Introduction to TURBOMOLE
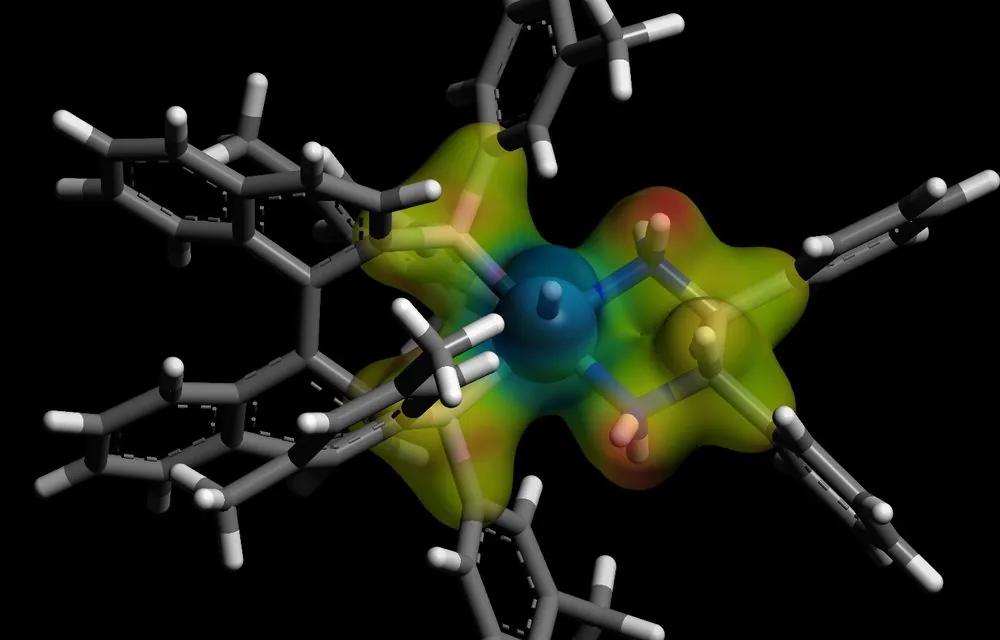
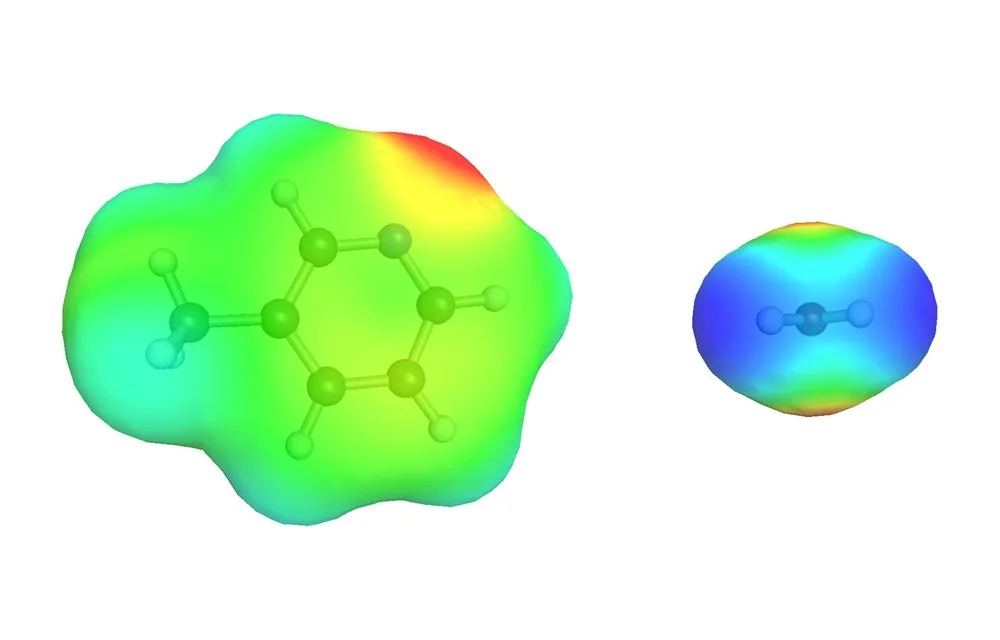
This course is an introduction to BIOVIA TURBOMOLE, the quantum chemistry program package for ab initio electronic structure calculations on molecules, clusters, periodic systems and solutions. Students learn to build or import structures and to set up, run and analyze calculations in TmoleX, the TURBOMOLE graphical user interface, and to choose between Hartree-Fock, density functional theory (DFT), MP2 and coupled cluster methods and suitable basis sets. The course covers structure optimization, transition state searches, vibrational frequencies with IR and Raman spectra, excited states and UV-Vis spectra with TDDFT, and solvent effects with COSMO, including the COSMO files used by COSMOtherm. Hands-on exercises are included.
- Level
- Fundamental
- Length
- 8 hours
- Product
- TURBOMOLE
- Delivery
- Instructor-led
Topics covered
- TURBOMOLE overview: methods, applications and program modules
- TmoleX interface: building, importing and pre-optimizing structures
- Methods: Hartree-Fock, DFT, MP2 and coupled cluster
- Basis sets, RI approximation and dispersion corrections
- Structure optimization and molecular symmetry
- Transition state searches and potential energy surface scans
- Vibrational frequencies, IR and Raman spectra
- Excited states and UV-Vis spectra with TDDFT
- Solvent effects with COSMO and COSMO files for COSMOtherm
- Running jobs on local or remote computers and analyzing results
Who it is for
Chemists and materials scientists in industry and academia who are new to TURBOMOLE, including those who run quantum chemical calculations only occasionally.